
Ready-to-Sequence Libraries. Flexible Sequencing Solutions.
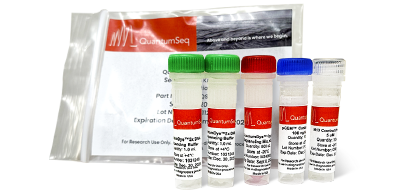
Quintara Sequencing Only provides a cost-effective solution for researchers who prepare their own sequencing libraries. We handle sequencing and demultiplexing, with flexible sequencing options to support a wide range of applications, including WGS, RNA-seq, amplicon sequencing, ATAC-seq, CUT&RUN, and single-cell sequencing. Get the sequencing data you need without the added cost of library preparation.


Flexible Options
Multiple Read Lengths & Data Outputs

Easy Submission
1000+ Free Drop Boxes

Reliable Quality Control
Library QC Before, Data QC After

More Savings
Capacity-Based Discounts
From library submission and pre-sequencing QC to flexible sequencing and data delivery, Quintara Seq-Only streamlines your sequencing workflow. Customize read length and data output, optionally add PhiX spike-in, and receive raw sequencing data with QC reports and optional downstream data analysis to support your research.

Submit Your Ready-to-Sequence Libraries

Quality Checks Before Sequencing

Flexible Read Lengths, Read Pairs & Optional PhiX

Raw Data & Optional Downstream Analysis
Read Length | Read Pairs | Price + |
2 x 75 bp | 20 M – 1000 M | Starting from $100 |
28 bp + 90 bp | 20 M – 1000 M | Starting from $100 |
2 x 150 bp | 20 M – 1000 M | Starting from $150 |
2 x 300 bp | 0.5 M – 400 M | Starting from $140 |
* Turnaround time may vary depending on sequencing capacity and scheduling. Contact us for the most accurate timeline.
+ Contact us to explore additional discounts and special pricing.
The i5 and i7 index sequences must each be 10 bp long. If your indexes are shorter than 10 bp, please add the appropriate number of padding bases from the adjacent primer sequence so that each index read totals 10 bp. Please confirm that your index sequences and padding bases are compatible with the sequencing configuration.
Yes. PhiX control consumes sequencing capacity and will reduce the number of reads allocated to your library. Please consider this when selecting your sequencing output. PhiX spike-in is optional and should be selected based on your library type and sequencing requirements.
One read pair = 2 reads. For paired-end sequencing, each read pair consists of one forward read (R1) and one reverse read (R2). For example, 20 M read pairs = 40 M total reads. The total number of bases also depends on the selected read length.
You will receive raw sequencing data in FASTQ format and applicable sequencing QC reports. Demultiplexed FASTQ files are provided when applicable. Optional downstream data analysis may also be available depending on your application and analysis requirements.

Bulk RNA sequencing platform by Quintara enabling transcriptome-wide studies.
Discover more
Quintara’s high-precision AmpExp platform supports targeted genomic studies.
Discover more
Rapid, affordable Nanopore sequencing from Quintara; plasmid DNA results in one day.
Discover more
We Go Fast, You Go Forward
 sales.us@quintarabio.com
 1-617-789-6111
 3563 Investment Blvd, #2, Hayward, CA 94545sales.us@quintarabio.com1-617-789-611165 Grove St, Suite 102, Watertown, MA 02472sales.us@quintarabio.com1-617-789-611110410 Corporate Dr. Suite 118, Sugar Land, TX 77478sales.us@quintarabio.com1-617-789-61116364 Ferris Square, Suite 102, San Diego, CA 92121
Quintara Biosciences © 2025 All Rights Reserved | Site Map | Terms of Use | Privacy Policy


 Login
 Order
 Contact Us
 Contact Us